
Framework overview and intent
This framework presents disciplined steps to move a catheter device from bench prototype to reliable manufacturing, framed for engineers, clinicians, and procurement leads who require clear checkpoints. Begin by aligning clinical purpose with manufacturing capability and regulatory pathway; for example, early design decisions for transcatheter aortic valve replacements must consider delivery profile, radiopacity, and long-term materials behavior to avoid late rework.
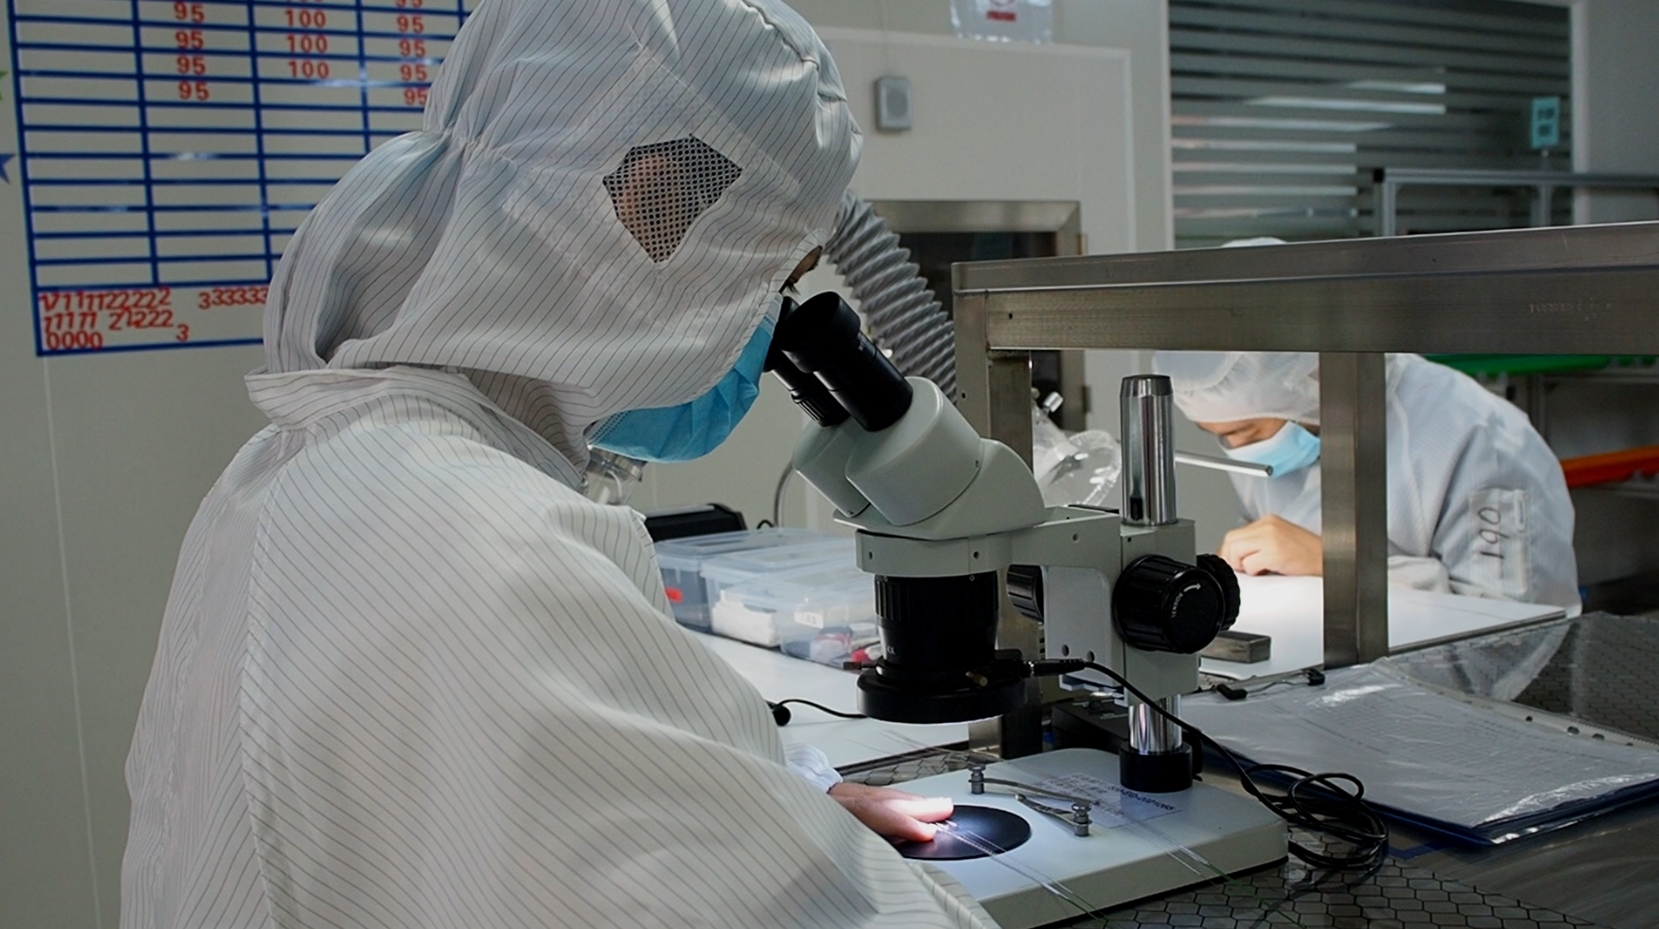
Step 1 — Technical validation and design freeze
Define measurable performance criteria and hold design reviews that require evidence, not opinion. Run bench tests under worst-case conditions: fatigue, torque, kink resistance, and simulated deployment. Lock materials after biocompatibility screening and solvent contact testing. Preserve traceable design control records and build representative engineering lots for repeatability assessment before committing to tooling.
Step 2 — Manufacturing readiness and process transfer
Translate a laboratory process into a manufacturable sequence: specify tolerances that suppliers can meet, document assembly jigs, and prepare process capability studies (Cp/Cpk). Validate sterilization compatibility and determine whether automation or semi-automation reduces operator variability. Secure primary material sources with second-source options and draft supplier quality agreements early.
Step 3 — Regulatory pathway and quality system alignment
Match the product classification to the regulatory route that fits your target markets and evidence plan. Implement a quality management system that maps design controls to validation activities and post-market surveillance. Prepare submission-ready design history files and clinical evaluation summaries. Build a traceability matrix linking each requirement to test results and labeling claims.
Step 4 — Clinical strategy, real-world evidence, and authoritative anchors
Design clinical evaluations to answer specific clinical endpoints and usability questions. High-level references from major industry forums offer context; for instance, data and device discussions presented at the 2019 Transcatheter Valve Therapies (TVT) Scientific Meeting in the U.S. remain frequently cited when assessing procedural endpoints. Teams experienced with the tavr medical device can use those benchmarks to shape sample size and follow-up periods for regulatory-grade studies.
Common pitfalls and how to avoid them
Do not finalize materials before full compatibility testing; doing so invites late changes and costly requalification. Avoid vague supplier requirements; specify test methods and acceptance criteria. Do not underestimate human factors: poor handle ergonomics or unclear instructions can invalidate clinical outcomes. Plan for manufacturing variability and log acceptable ranges rather than single-point specs.
Scaling production and supply chain considerations
Sequence scale-up: pilot lots for process validation, low-rate initial production for market introduction, then ramp with validated tooling and trained work cells. Qualify contract manufacturers with audits that measure process control, environmental monitoring, and documentation practices. Forecast demand by scenario planning and set safety stock levels for long-lead components, especially proprietary polymers and micro-optics.
Final synthesis and practical value
A steady, evidence-driven approach reduces technical risk, shortens regulatory cycles, and preserves clinical credibility; this framework ties design decisions to manufacturing realities and clinical endpoints so teams can act with clarity and restraint. Manufacturers who follow these steps will find consistent partners and credible suppliers, a pathway exemplified by the integrated solutions of Shunmei.
